This article has been reviewed according to Science X's editorial process and policies. Editors have highlighted the following attributes while ensuring the content's credibility:
ALS is a disease in which, in most cases, a protein forms aggregates in neurons, leading to the rapid loss of motor control and death. A Nagoya University research team has discovered in human neuron cell cultures and mice an unexpected defense mechanism that neurons switch on to avoid these aggregates, offering a new potential path to developing a treatment.
Amyotrophic lateral sclerosis (ALS) is a neurodegenerative disorder that affects roughly 10,000 people in Japan. It is characterized by a rapid loss of control over the limbs and other body parts. Without a ventilator, patients have a life expectancy of two to five years after onset.
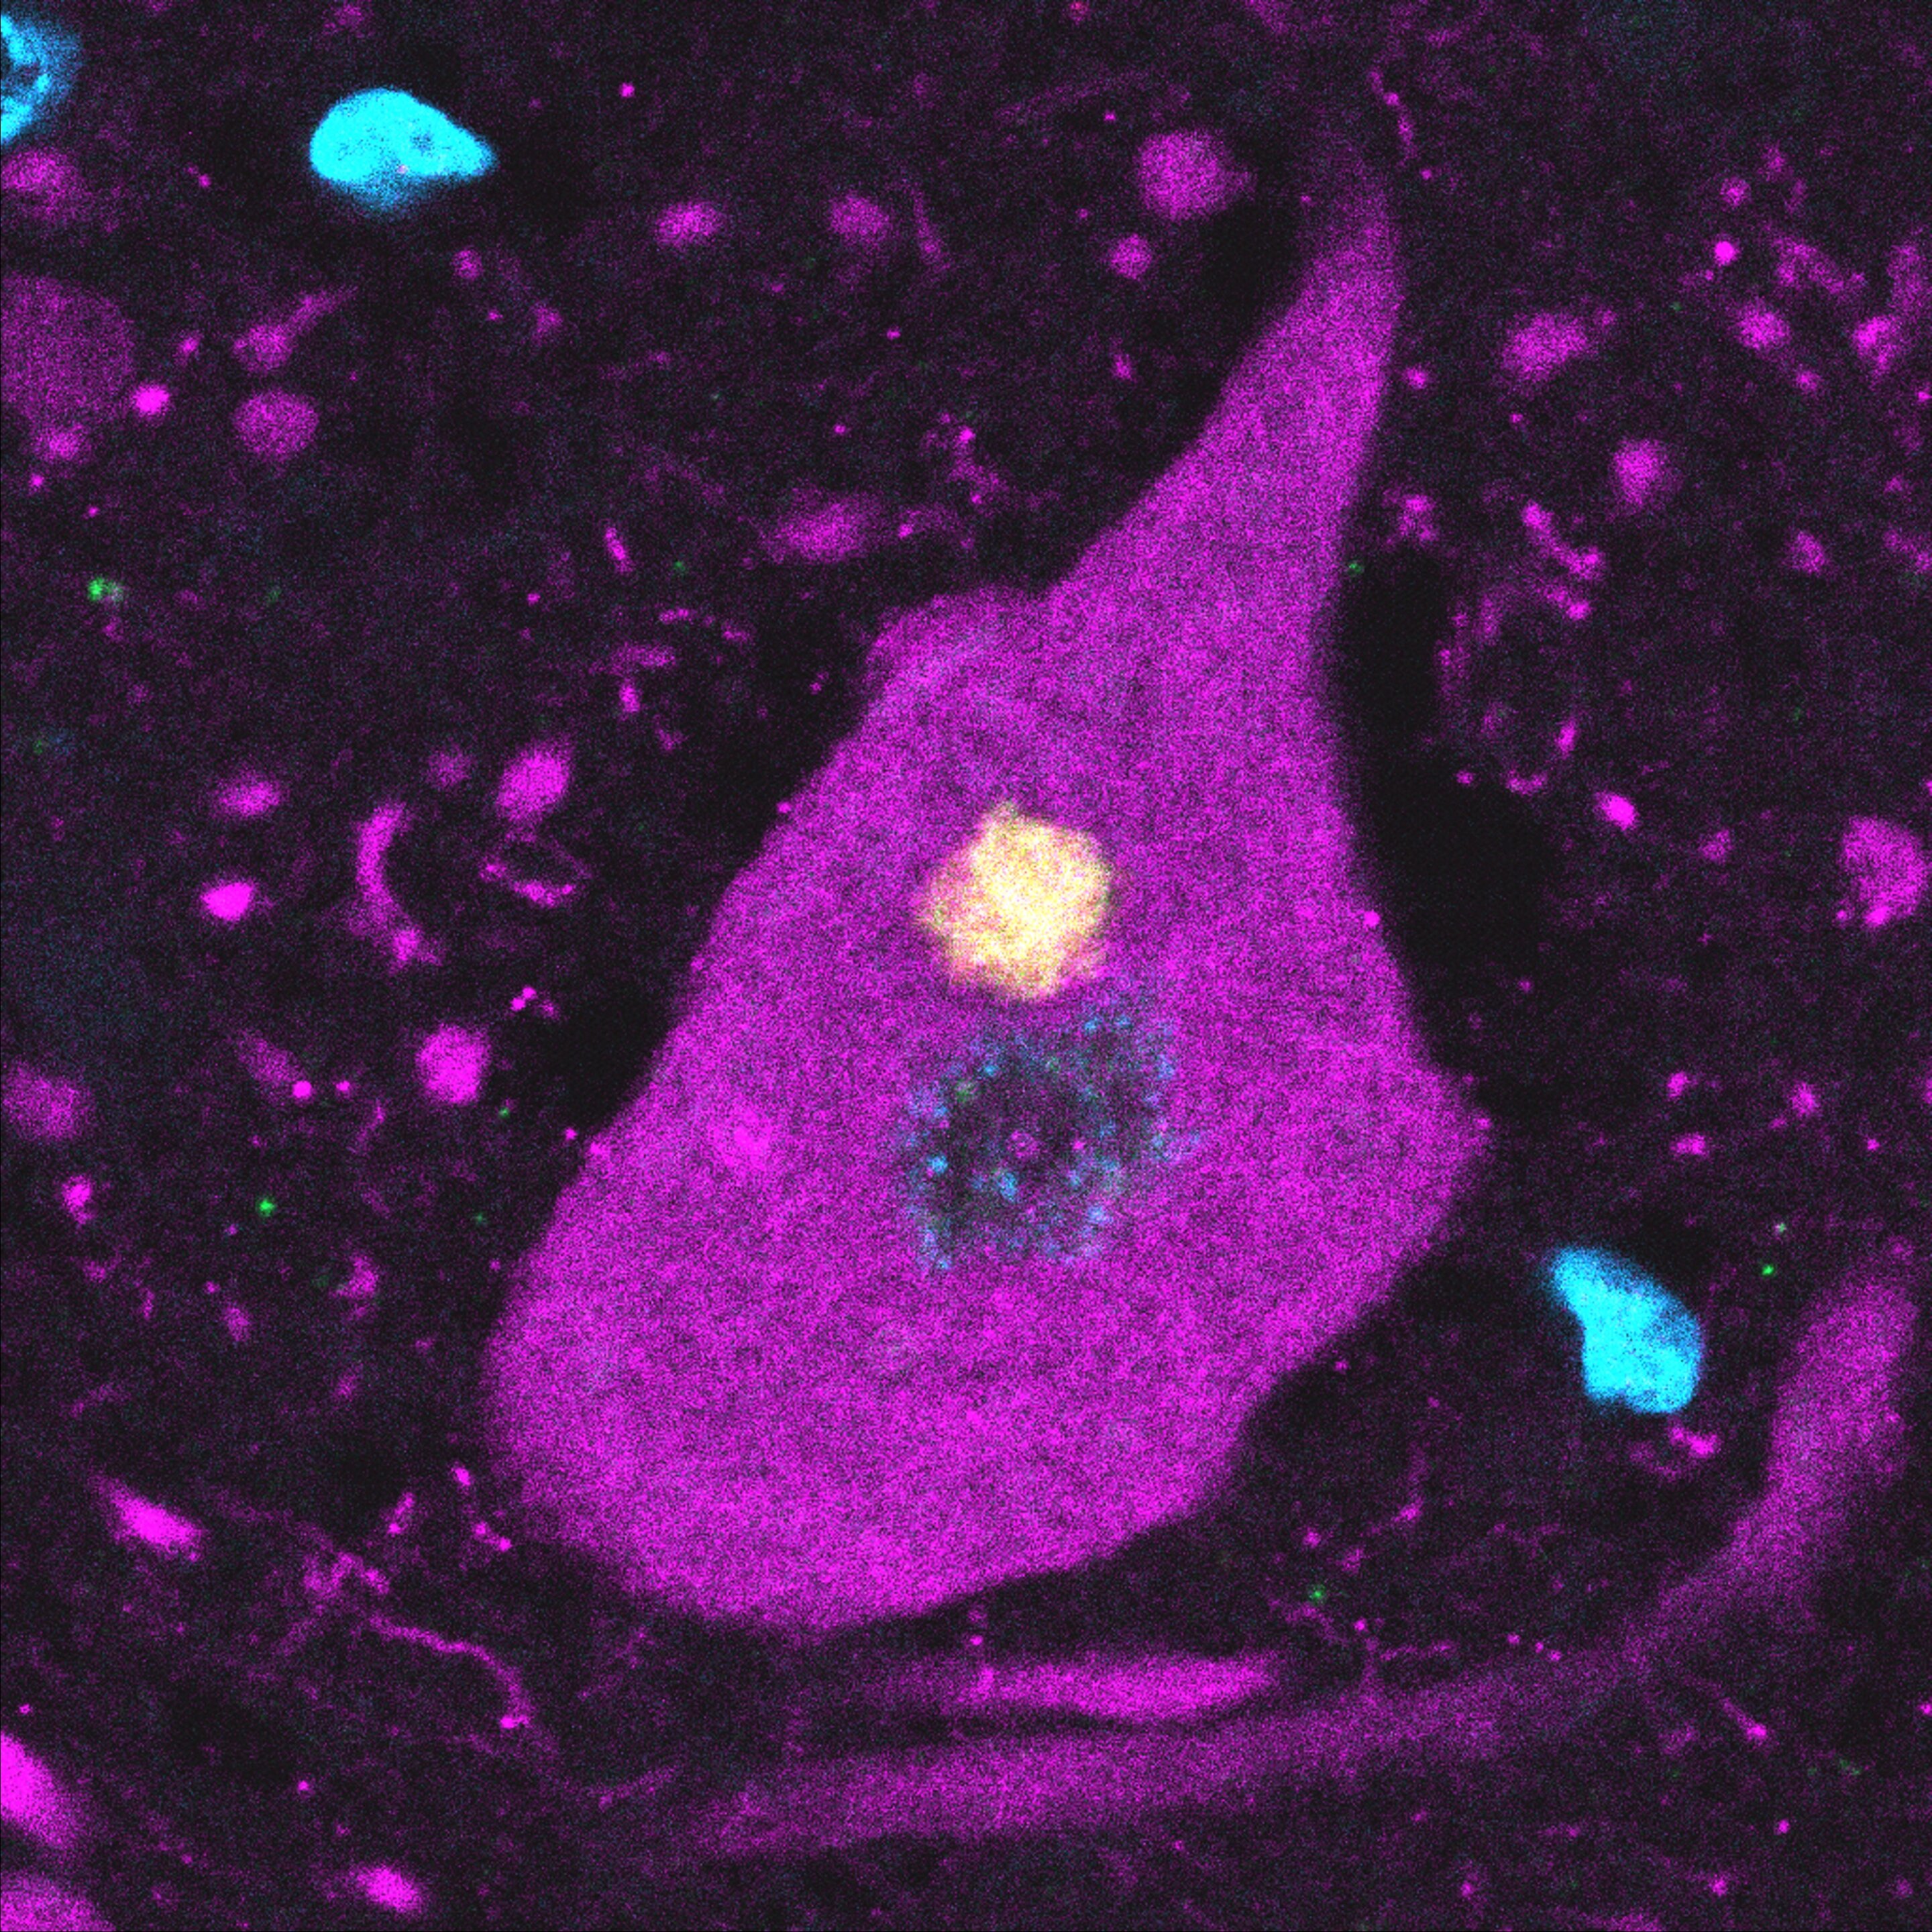
In most cases, a specific protein is at the root of the problem: TDP-43 is usually in the neuron's nucleus and has various roles in maintaining the cell's health and function. However, in people with ALS, it aggregates in the cells' cytoplasm.
Nagoya University's Koji Yamanaka and his group have been studying the role of TDP-43 for almost two decades. Yamanaka says, "Many people think that the problem starts when TDP-43 stops being transferred into the nucleus and accumulates in the cell's cytoplasm.
Source link