
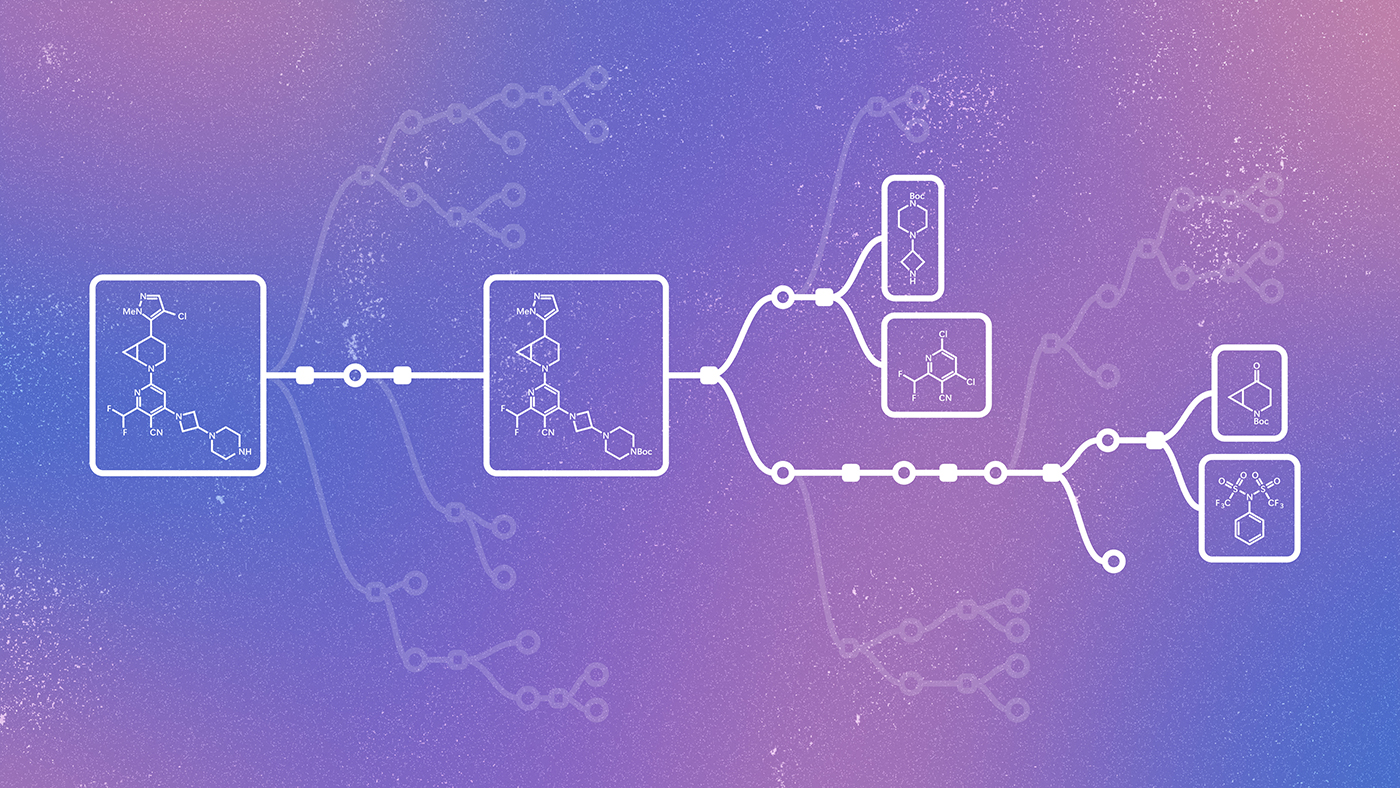
Developing new medicines and materials requires making new molecules but planning how to make them is still largely manual, time-consuming, and costly. RetroChimera automatically proposes high-quality synthesis routes. The model combines two strong models with complementary strengths, learning how to rank their proposals to produce better predictions than either alone. In blind tests, PhD-level chemists prefer RetroChimera’s individual reaction predictions over preceding models and recorded literature reactions.
Custom-made molecules are unlocking advances in modern medicine, smart materials, and sustainable agriculture. Yet, progress is slowed by chemical synthesis—the time-consuming process of making new molecules from simpler building blocks in the lab. In addition, synthesis is a significant driver of drug development costs. So even as computational methods make it possible to explore large numbers of novel molecules, finding practical ways to synthesize them remains a critical challenge.
Retrosynthesis approaches this problem by working backwards from a target molecule, breaking it down step by step into simpler precursors (Figure 1). This process is comparable to playing strategic board games like chess and Go. It involves contemplating a wide range of possible immediate moves, or individual disconnections, while also requiring high-level strategic thinking to reach the end-to-end synthesis plan. However, the number of possible moves in retrosynthesis is much larger than in board games, and it is not obvious which moves would be available for a given molecule. Existing systems face major challenges, including recalling rare but strategically important reactions, robustness beyond the training distribution, and aligning with chemists’ expectations. As a result, retrosynthesis often requires highly specialized expertise, which hinders scaling and automation of scientific discovery.
Source link





